Summary:
FDA warning letter recovery is the structured process a company follows after receiving a warning letter — investigating root causes, submitting a written response within 15 business days, implementing verified corrective actions, and passing a follow-up inspection to reach FDA close-out. Full recovery typically takes 18–36 months for pharmaceutical manufacturers.
If your facility has received a a warning letter, or you want to be ready before one arrives, this guide walks through the entire journey: how a routine inspection turns into a warning letter, what recovery actually requires, and why some companies close out in 12 months while others are still fighting the same citation three years later.
The Complete FDA Enforcement Journey
Recovery only makes sense in the context of the full enforcement lifecycle. Each stage narrows your options and raises the cost of getting it wrong.
| Stage | What Happens | Typical Duration |
|---|---|---|
| 1. Inspection | An FDA investigator conducts an on-site or remote CGMP inspection under the FD&C Act | 1–2 weeks |
| 2. Form 483 | Investigator issues a list of objectionable conditions observed during the inspection | Issued at inspection close-out |
| 3. Company response to 483 | Company submits a written corrective action plan | Due within 15 business days |
| 4. Warning Letter | Issued if the FDA finds the 483 response inadequate, or if violations are significant enough on their own | Weeks to months after the 483 |
| 5. Remediation / recovery | Company investigates, implements, and documents corrective and preventive actions | 6–24 months |
| 6. Reinspection | FDA verifies implementation, often unannounced | Scheduled once remediation evidence is submitted |
| 7. Close-out | FDA issues a close-out letter confirming the violations have been adequately addressed | 18–36 months from the original warning letter, on average |
A Form 483 and a warning letter are often confused. A Form 483 lists observations made during a single inspection — it is not itself an enforcement action. A warning letter is the FDA’s formal notice that your company’s prior response failed to correct the problem, or that the violations were serious enough to skip straight to a letter. Warning letters are public, are posted on FDA.gov, and can trigger import alerts, injunctions, or consent decrees if unresolved — which is why treating the 483 stage seriously is the cheapest form of warning letter prevention.
Why Warning Letters Are Increasing
Recovery plans built on outdated assumptions fail. Here’s what current enforcement data shows:
- Volume is up sharply. FDA issued 303 drug and biologics warning letters in FY2025, up from 190 in FY2024 — both 483s and warning letters are at five-year highs.
- The top citation is procedural, not technical. The most cited Form 483 observation is failure to follow written quality unit procedures under 21 CFR 211.22(d), cited 184 times in FY2024 — a governance failure, not a lab error.
- Laboratory controls and OOS handling dominate. Out-of-specification investigation inadequacies remain the single most cited pharmaceutical issue, with FDA investigators increasingly testing whether root cause conclusions are scientifically defensible rather than paperwork exercises.
- Data integrity is a multi-year focus. Falsified chromatography data, audit trail manipulation, and shared login credentials continue to draw scrutiny, especially at foreign manufacturing sites.
- Contract manufacturers are squarely in scope. FDA has made clear that CMOs and contract labs are treated as extensions of the brand owner’s own facility — outsourcing does not transfer accountability.
- Recovery takes longer than most companies plan for. Published FDA close-out data puts average resolution at 18–36 months for pharmaceutical manufacturers, which is why a rushed, single-cycle response so often backfires.
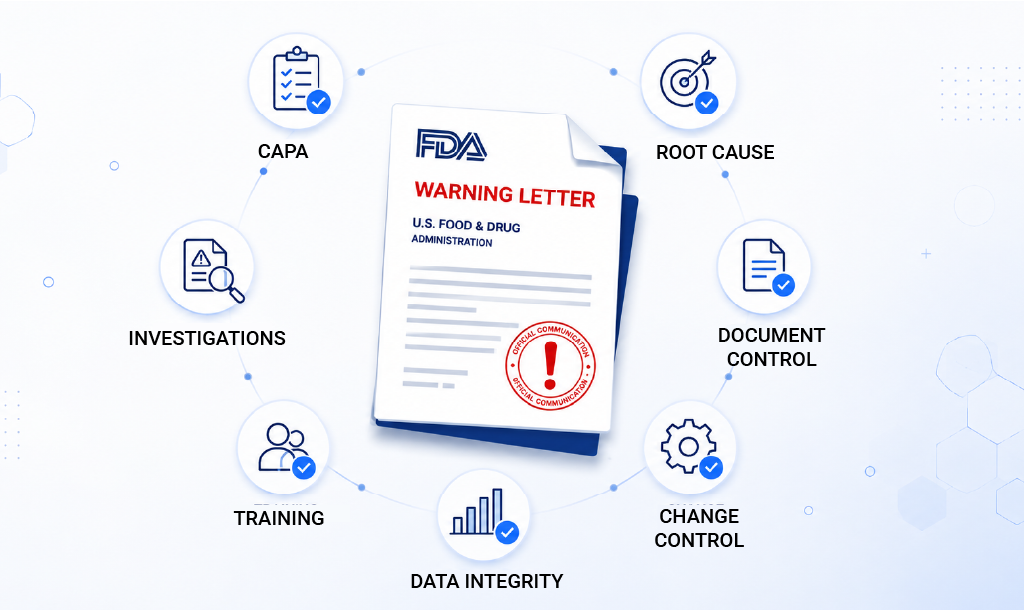
This matches a pattern we found reviewing 148 FDA warning letters directly: the same CAPA, investigation, training, and change control failures repeat across companies, industries, and years. See our full analysis of 148 FDA warning letters and the quality system patterns behind them for the underlying data.
The 8-Step FDA Warning Letter Recovery Framework
This is the sequence that separates a one-cycle recovery from a multi-year enforcement relationship. Each step names who typically owns it.
Step 1 — Stabilize in the first 48 hours (Executive + Plant Manager)
Do not respond reactively. Notify executive leadership and legal counsel immediately, preserve all relevant records, and resist the urge to make verbal commitments to the FDA before your team has assessed the full scope. If distributed product may be affected, begin a risk assessment for market action in parallel — the FDA specifically looks for this in later reviews.
Step 2 — Assemble a cross-functional response team (Quality Manager leads)
A warning letter is never purely a quality problem. Build a team spanning Quality, Regulatory Affairs, Manufacturing/Plant Operations, Legal, and an executive sponsor. Each function owns a distinct piece: Quality owns investigation rigor, Regulatory owns FDA communication and submission format, Plant Managers own operational feasibility of proposed fixes, and executives own resourcing and accountability.
Step 3 — Map every observation line by line (Regulatory Affairs)
Break the letter into individually numbered observations. For each one, capture: the exact regulatory citation, the FDA’s stated concern, related historical 483s or deviations, and the business unit responsible. Untangling combined observations at this stage prevents partial responses later.
Step 4 — Conduct root cause analysis — not root cause labeling (Quality Manager)
This is where most recoveries stall. “Human error” and “equipment malfunction” are conclusions the FDA routinely rejects unless you can show why the system allowed the error to occur. Effective analysis pulls training records, maintenance logs, and prior deviations — not just the immediate incident file. Our review of warning letter patterns found root cause failures cited alongside CAPA and investigation failures far more often than in isolation, which is why they need to be solved together, not sequentially.
Step 5 — Build a CAPA plan the FDA can verify (Quality Manager + Plant Manager)
Every corrective action needs four things: a documented root cause, an owner, a due date, and a defined effectiveness check. A CAPA that closes without evidence the problem stopped recurring is one of the most common reasons the FDA escalates rather than closes a case. If your facility’s gaps trace back to CGMP fundamentals — batch release testing, supplier qualification, process validation, or stability programs — see our breakdown of four CGMP violations that led to an FDA import ban for how each one maps to a specific prevention control.
Step 6 — Draft and submit the written response within 15 business days (Regulatory Affairs, reviewed by Executive + Legal)
The 15-business-day clock is non-negotiable. If full remediation cannot realistically finish in that window — and for systemic issues, it usually can’t — the response should still demonstrate immediate containment actions, a committed remediation timeline with milestones, and evidence the investigation is already underway. A response of “we will investigate” without specifics is treated as inadequate.
Step 7 — Implement corrective actions and document as you go (Plant Manager + Quality Manager)
Recovery is verified on paper as much as on the floor. Every implemented action needs a traceable record: training completed, procedures revised and re-approved, equipment requalified, and — critically — effectiveness checks scheduled and completed on time. Companies that centralize this evidence recover faster than those reconstructing it from spreadsheets and email at reinspection time.
Step 8 — Prepare for reinspection and request close-out (Executive + Regulatory Affairs)
Once major CAPAs are implemented and verified, submit updated evidence to the FDA and prepare the site for an announced or unannounced reinspection. A close-out letter is only issued once the FDA independently confirms the fixes hold — self-reported completion is not sufficient.
Recovery Timeline: What to Expect, Month by Month
| Timeframe | Milestone |
|---|---|
| Day 0–15 (business days) | Written response submitted to FDA |
| Weeks 2–8 | Root cause investigations completed; immediate containment actions in place |
| Months 2–6 | CAPA implementation across affected processes; interim status updates if FDA requests them |
| Months 6–12 | Effectiveness checks completed; internal mock inspection conducted |
| Months 12–24 | FDA reinspection scheduled and completed |
| Months 18–36 | Close-out letter issued (average range across pharmaceutical manufacturers) |
Timelines compress significantly for companies with strong evidence trails going in, and extend when CAPAs are reopened due to recurrence — which, per FDA enforcement patterns, happens most often when the original root cause analysis addressed the symptom rather than the system.
Recovery Checklist & CAPA Response Template
Use this as a working checklist for your response team. It’s built to be copied into your own tracker or CAPA software.
Immediate Actions (Week 1)
- Confirm receipt and log the 15-business-day response deadline
- Notify executive sponsor, legal counsel, and affected site leadership
- Freeze and preserve all related batch, deviation, and training records
- Assemble the cross-functional response team with named owners
- Begin a risk assessment for any product already in distribution
Investigation Phase (Weeks 1–4)
- Number and categorize every observation in the letter
- Pull related historical 483s, deviations, and prior CAPAs for pattern review
- Complete root cause analysis for each observation category
- Identify whether observations are isolated or systemic across sites/products
Response Drafting (Weeks 2–3)
- Draft observation-by-observation response with committed timelines
- Include evidence of containment actions already taken
- Legal and executive review before submission
- Submit via the method and address specified in the letter
CAPA Template (per observation)
- Observation reference and regulatory citation
- Root cause (systemic, not symptomatic)
- Corrective action and responsible owner
- Preventive action to stop recurrence across similar products/processes
- Due date and interim milestones
- Effectiveness check method and verification date
- Supporting evidence/document links
Pre-Reinspection Readiness (Ongoing)
- Internal mock inspection against all cited observations
- Confirm every CAPA has closed with documented effectiveness evidence
- Refresh training records tied to any procedure changes
- Compile a centralized, inspector-ready evidence package
Case Examples: Why Recoveries Succeed or Fail
The following are anonymized composites based on publicly documented enforcement patterns, not individual companies.
Case A — Recovery in one cycle. A mid-size sterile manufacturer received a warning letter citing inadequate OOS investigations. Rather than responding observation-by-observation in isolation, the quality team traced the pattern back to a single root cause: lab analysts were authorized to retest without a documented, pre-approved retest protocol. The CAPA redesigned the OOS workflow itself, retrained the full lab staff, and built an effectiveness check into the next four quarterly trend reviews. FDA closed the letter after one reinspection, roughly 14 months after issuance — faster than the average, because the response addressed the system, not just the cited batches.
Case B — Recovery that stalled. contract manufacturer received a warning letter for cross-contamination risk tied to cleaning validation gaps. The initial response focused narrowly on the specific equipment named in the letter, added supplemental cleaning steps, and closed the CAPA without a formal effectiveness study. Eighteen months later, a similar finding recurred on a different product line using shared equipment — because the underlying cleaning validation program itself was never rebuilt. The case remained open past the 24-month mark, and the FDA’s follow-up correspondence noted the recurrence explicitly as evidence of an ineffective original response. The lesson mirrors what shows up across FDA enforcement broadly: it’s rarely one catastrophic event — usually just a handful of gaps in quality processes that any manufacturer could have, and most don’t catch until it’s too late, as we detailed in our review of four CGMP violations behind a recent FDA import ban.
Recovery Mistakes by Role
Quality Managers most often fail by treating each observation as a standalone fix rather than checking for the same root cause elsewhere in the quality system — the FDA frequently finds the identical gap on a different product line during reinspection.
Regulatory Affairs sometimes submit a response that reads as legally cautious but operationally vague — committing to “review and enhance” procedures without naming specific actions, owners, and dates. The FDA treats this as evasive rather than reassuring.
Plant Managers under-resource the operational side of CAPA implementation, prioritizing production continuity over corrective action deadlines — which shows up later as overdue, unescalated CAPAs during reinspection.
Executives often make the mistake of treating the warning letter as a Quality department problem to delegate fully, rather than a business continuity risk requiring cross-functional resourcing and visible leadership accountability — something FDA investigators explicitly assess through interviews during reinspection.
How a Connected QMS Shortens Recovery
Every mistake above traces back to the same structural issue: information about deviations, CAPAs, training, and document control living in disconnected systems. That’s the exact pattern our review of 148 warning letters found — CAPA problems show up next to investigation problems, investigation problems show up next to root-cause problems, and training gaps show up next to document control gaps.
BizPortals QCFlow is built to close those gaps during recovery, not just after it:
- CAPAs cannot close without documented effectiveness verification
- Investigations are pre-linked to related deviations, prior CAPAs, and training history
- Document approvals automatically trigger training assignments — no manual handoff to miss
- Real-time dashboards replace the multi-week manual evidence compilation that slows reinspection prep
If your recovery plan depends on pulling data from spreadsheets, shared drives, and email threads, a QMS review is worth doing before your next FDA interaction. Book a demo to see how QCFlow maps to your open observations.
FAQs
-
What is FDA warning letter recovery?
FDA warning letter recovery is the process of investigating and correcting the violations cited in a warning letter, submitting a written response within 15 business days, implementing and verifying corrective actions, and passing a follow-up FDA inspection to receive a close-out letter.
-
How long does FDA warning letter recovery take?
On average, 18–36 months from issuance to close-out for pharmaceutical manufacturers, though single-cycle recoveries with strong root cause analysis and documentation can close in 12–14 months.
-
What is the deadline to respond to an FDA warning letter?
15 business days from the date of the letter. Even if full remediation will take longer, the response must be submitted within this window and should include containment actions and a committed timeline.
-
What’s the difference between a Form 483 and a warning letter?
A Form 483 lists observations from a single inspection and is not itself an enforcement action. A warning letter is issued when the FDA finds the 483 response inadequate or the violations serious enough on their own, and it is a formal, public enforcement action.
-
Can a company request a meeting with the FDA during recovery?
Yes, companies can request a meeting to discuss their recovery plan, but the FDA does not use these meetings to provide corrective guidance — the burden remains on the company to design and defend its own remediation plan.
-
Why do some companies receive repeat warning letters for the same issue?
Repeat citations happen when a CAPA addresses the symptom named in the letter without fixing the systemic cause — the same failure type then resurfaces on a different product, line, or site.
-
What should a warning letter response include?
An observation-by-observation breakdown, documented root cause for each, specific corrective and preventive actions with owners and dates, evidence of any immediate containment already taken, and a realistic implementation timeline.
-
Does a warning letter affect a company’s ability to sell products?
Not automatically, but unresolved warning letters can lead to import alerts on foreign facilities, injunctions, or consent decrees, and they are publicly posted on FDA.gov, which affects customer and partner due diligence.
-
What role does an executive sponsor play in recovery?
Executives are responsible for resourcing the cross-functional response team, removing operational obstacles to CAPA deadlines, and demonstrating leadership accountability — something FDA investigators assess directly during reinspection interviews.